Tucidinostat (Chidamide) Tucidinostat HDAC target inhibition synergizes with Bortezomib in gastric cancer
Copy code to add this image to your page
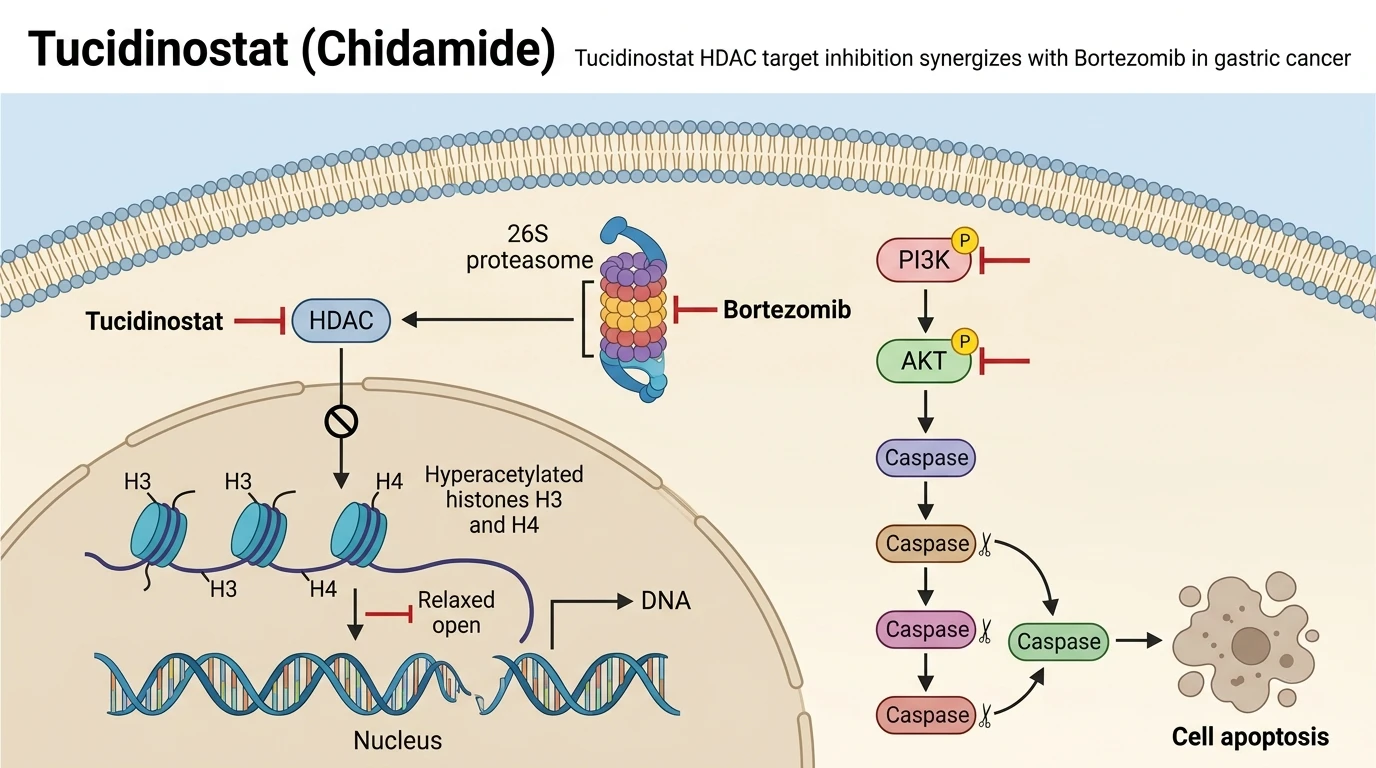
Gastric carcinogenesis involves complex driver mechanisms, prominent among which is the aberrant modification of chromatin structures through histone deacetylase hyperactivation alongside constitutively active intracellular survival cascades. Selective enzyme inhibition offers a directed therapeutic approach. Tucidinostat (Chidamide) functions as a benzamide-class, orally bioavailable selective inhibitor targeting Class I (HDAC1, HDAC2, HDAC3) and Class IIb (HDAC10) enzymes. By binding the zinc-dependent catalytic pocket of these deacetylases, Tucidinostat (Chidamide) blocks core histone deacetylation, resulting in sustained hyperacetylation of core histone tails H3 and H4. This chromatin remodeling enables re-expression of silenced tumor-suppressor genes and alters downstream signaling cascades vital for tumor cell survival.
In gastric cancer cell models, specifically MGC-803 and BGC-823 lines, monotherapy with HDAC inhibitors frequently encounters compensatory survival signals. Epigenetic modulation through Tucidinostat HDAC targeted action interacts directly with proteasomal degradation pathways. The 26S proteasome inhibitor Bortezomib, when administered in low-dose combination with Tucidinostat (Chidamide), produces a marked synergistic effect on cell proliferation, migration, and invasive capability under tested in vitro assay conditions. This biological outcome stems from reciprocal signaling cross-talk suppression. While individual agents exert modest responses at reduced concentrations, their co-administration induces pronounced cellular apoptosis.
At the molecular node level, the combined regimen modulates the phosphoinositide 3-kinase (PI3K)/Akt axis, a primary driver of survival and metastatic dissemination in gastric malignancies. Under experimental treatment conditions, Bortezomib potentiates the HDAC-inhibitory capacity of Tucidinostat (Chidamide). This functional synergy elevates global histone H3 and H4 acetylation levels beyond those achieved with monotherapy. Concurrently, western blot analyses reveal a substantial decline in phosphorylated PI3K (p-PI3K) and phosphorylated AKT (p-AKT) levels, demonstrating that epigenetic enzyme suppression interacts directly with receptor-downstream kinase cascades. Downstream effector suppression destabilizes cell survival signals, initiating caspase cleavage cascades and nuclear fragmentation.
Phenotypic assays corroborate these molecular alterations across tested gastric cell populations. Transwell and wound-healing assays demonstrate that low nanomolar co-treatments attenuate cell migration and invasion capabilities. Cellular proliferation metrics drop consistently in dose-response matrices, confirming true synergistic interaction as quantified by combination index calculations. These observations illustrate how dual inhibition of proteasomal processing and histone modification dismantles survival networks in aggressive gastric phenotypes. Investigating these epigenetic regulatory mechanisms provides essential insights into targeted combination regimes.
In vivo translation of this strategy was verified using BALB/c nude mouse xenograft models bearing MGC-803 subcutaneous tumors. Subcutaneous tumor burden was significantly restricted in animals receiving low-dose combined therapy compared to vehicle or single-agent controls over the experimental timeline. Systemic safety profiles remained intact. Hepatorenal biomarker panels, including serum alanine aminotransferase, aspartate aminotransferase, blood urea nitrogen, and creatinine levels, exhibited no statistically significant elevation above baseline controls. Body weight trajectories remained stable throughout the administration period. These empirical observations establish that combined proteasome and selective HDAC inhibition delivers potent antitumor efficacy without inducing prohibitive systemic toxicity within tested murine models.
Synergistic Epigenetic and Proteasomal Inhibition Dynamics
Copy code to add this image to your page
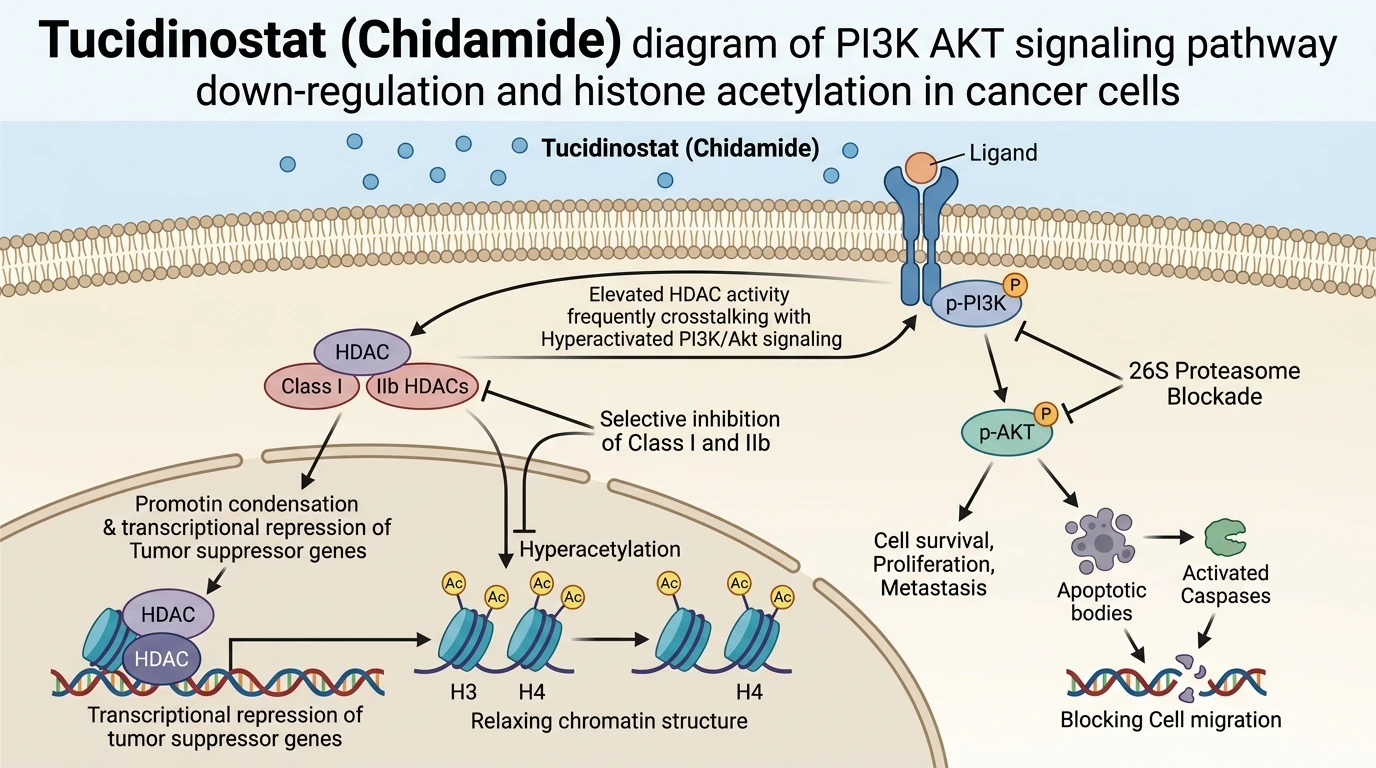
Tucidinostat (Chidamide) selectively inhibits Class I and IIb histone deacetylases, promoting core histone H3/H4 acetylation and epigenetic gene reactivation. When combined with the 26S proteasome inhibitor Bortezomib, low-dose administration produces synergistic antitumor activity in gastric cancer models. Mechanistically, Bortezomib potentiates HDAC inhibition, suppressing phosphorylated PI3K and AKT levels while downregulating cell proliferation, migration, and invasion. This dual strategy induces intrinsic mitochondrial apoptosis in MGC-803 and BGC-823 cells. In vivo xenograft experiments demonstrate marked tumor growth inhibition without hepatorenal toxicity.
Potentiation of Histone Acetylation and PI3K/Akt Suppression
Dual targeting of core histone deacetylases and the proteasome alters chromatin dynamics and survival kinase signaling. In MGC-803 and BGC-823 gastric carcinoma cells, co-treatment with low-dose Bortezomib amplifies Tucidinostat HDAC inhibition. Histone H3 and H4 acetylation levels increased significantly. Signal transduction through the PI3K/Akt pathway decreased in a dose-dependent manner. Phosphorylated AKT levels dropped markedly within hours of exposure. Downstream signaling ceased. Mechanistically, proteasome inhibition prevents the degradation of key negative regulators of kinase pathways, while Class I and IIb HDAC blockade alters transcriptional output. This dual action destabilizes oncogenic signaling networks. Cellular viability dropped sharply across treated cohorts. Importantly, these biochemical changes occurred at sub-cytotoxic individual concentrations, confirming strong pharmacological synergy in vitro.
Attenuation of Gastric Cancer Migration and Apoptosis Induction
Cellular functional assays confirm distinct phenotypic changes following combination treatment. In vitro transwell and wound-healing assays revealed reduced cellular migration. Matrix invasion decreased substantially under co-treatment conditions. Apoptotic rate rose rapidly. Cleaved caspase-3 and cleaved PARP levels increased, marking irreversible progression toward cell death. The selectivity margin remained intact. Quantitative combination index metrics demonstrated true synergism across multiple dosage ratios. Low-dose regimens achieved superior efficacy compared to high-dose single-agent therapies. Proliferation stopped rapidly. These observations confirm that dual epigenetic and proteasomal blockade overrides anti-apoptotic defenses in refractory gastric cancer lines.
In Vivo Tumor Suppression and Hepatorenal Tolerability Profile
In vivo validation confirmed the therapeutic potential of this dual strategy within murine models. In BALB/c nude mice bearing MGC-803 xenografts, low-dose combination therapy restricted tumor growth significantly. Tumor volume expansion halted over the 21-day observation period. Monotherapy cohorts exhibited persistent tumor expansion. Histological examination showed increased intra-tumoral necrosis and elevated apoptotic indices. Systemic toxicity remained minimal. Hepatorenal biomarkers—including serum ALT, AST, blood urea nitrogen, and creatinine—remained within normal physiological ranges. Animal body weights remained constant throughout the dosing schedule. These findings demonstrate that combined low-dose administration achieves robust in vivo tumor control without inducing cumulative visceral toxicity.