Neratinib from Selleck Covalently Inhibits HER2, EGFR, and HER4 in Metastatic Breast Cancer
Copy code to add this image to your page
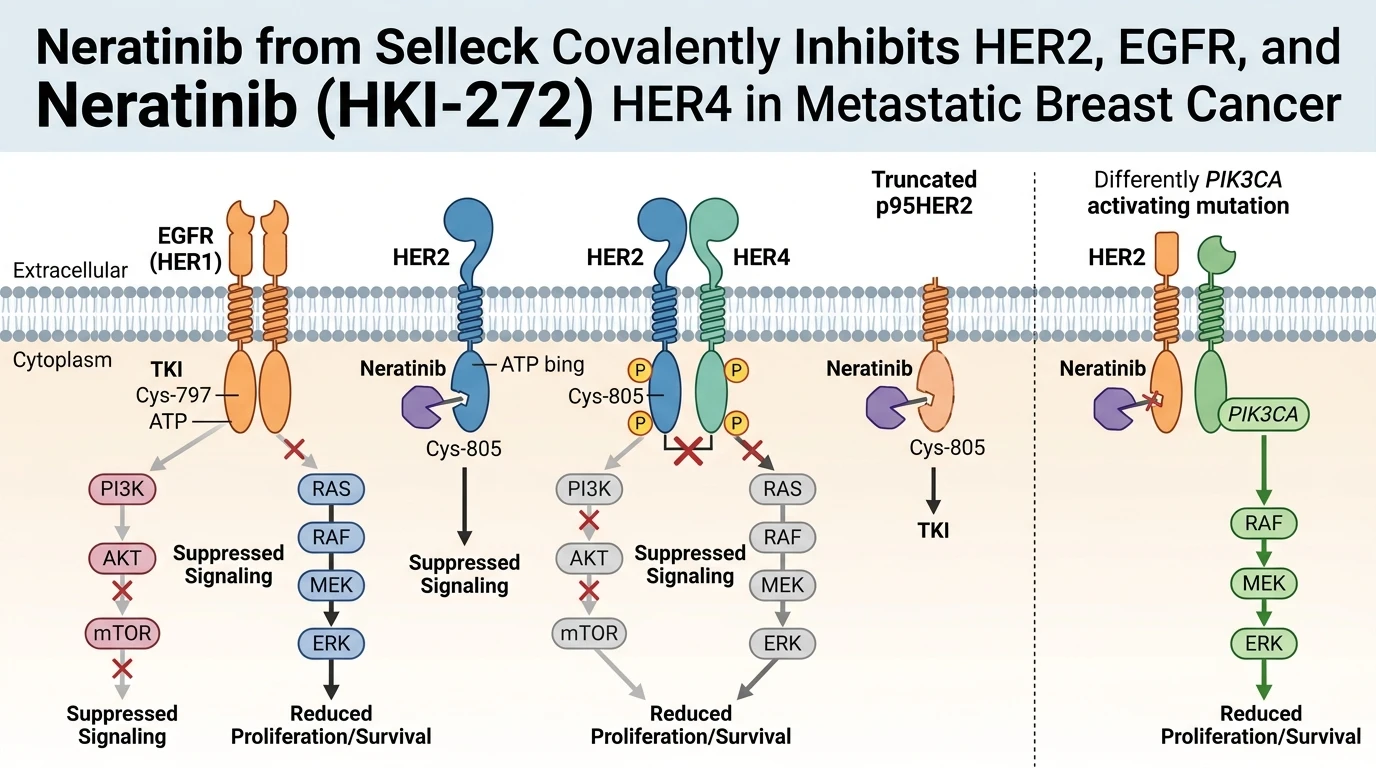
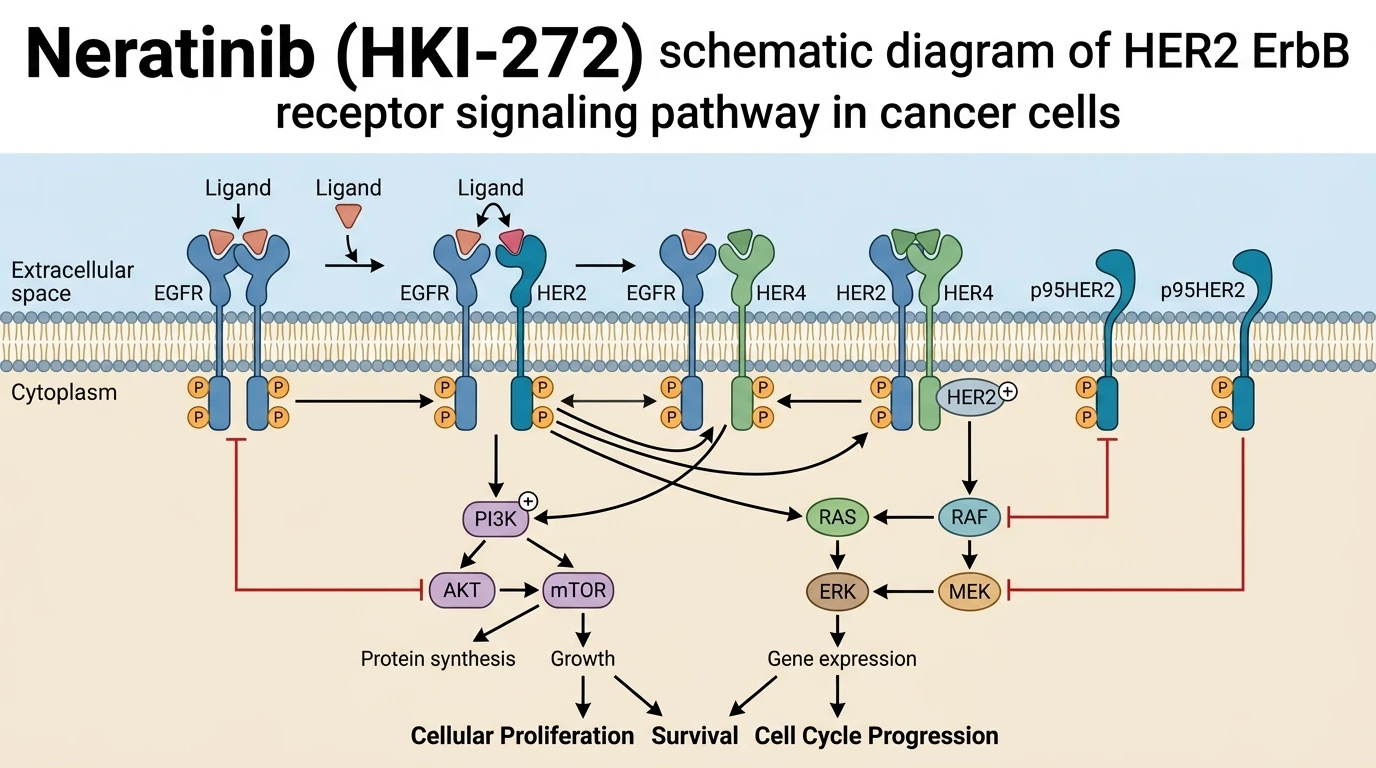
Targeted therapeutic strategies in HER2-positive metastatic breast cancer have evolved beyond initial monoclonal antibody approaches to encompass small-molecule tyrosine kinase inhibitors capable of targeting intracellular catalytic domains. Neratinib (HKI-272) is an orally administered, irreversible pan-HER tyrosine kinase inhibitor that targets human epidermal growth factor receptor 2 (HER2/ERBB2), epidermal growth factor receptor (EGFR/HER1), and HER4 (ERBB4). Unlike reversible inhibitors such as lapatinib, Neratinib from Selleck forms a permanent, covalent bond with a conserved cysteine residue within the ATP-binding pocket of these receptors—specifically Cys-805 in HER2 and Cys-797 in EGFR. This covalent adduct prevents ATP binding and irreversibly blocks transphosphorylation, resulting in sustained suppression of downstream signaling pathways including the phosphatidylinositol 3-kinase (PI3K)-AKT axis and the mitogen-activated protein kinase (MAPK) cascade.
The phase III NALA clinical trial provided a platform to evaluate the efficacy of neratinib in combination with capecitabine (N+C) against lapatinib plus capecitabine (L+C) in patients with HER2-positive metastatic breast cancer who had received two or more prior HER2-directed regimens. A comprehensive biomarker sub-analysis of pre-treatment tumor specimens from this cohort sought to clarify molecular correlates of progression-free survival (PFS) and refine patient stratification criteria for pan-HER TKI therapy.
Quantitative evaluation of HER2 protein expression across patient samples demonstrated that higher receptor density strongly correlates with enhanced clinical benefit from N+C relative to L+C. Protein expression levels were determined using multiple assay methodologies, including standard immunohistochemistry (IHC), semi-quantitative H-scoring, and the quantitative HERmark fluorescence assay. Patients whose tumors displayed elevated HER2 protein expression experienced superior PFS under N+C treatment. Mechanistically, high receptor density reflects strong oncogenic addiction to ErbB signaling, rendering tumor cells acutely dependent on receptor kinase activity and therefore highly responsive to irreversible catalytic inhibition.
This predictive association between elevated HER2 expression and extended PFS was most pronounced in hormone receptor-negative (HR-) metastatic breast cancer. In HR-negative tumors, the absence of crosstalk between estrogen or progesterone receptor signaling and the ErbB network means cell survival relies almost entirely on HER2-driven cascades. Consequently, complete and irreversible blockade of HER2 catalytic activity exerts a maximal anti-proliferative impact in this subpopulation, whereas hormone receptor-positive tumors may utilize alternative steroid signaling axes to sustain cell survival under kinase inhibitor pressure.
The biomarker analysis further illuminated the impact of structural receptor variants, specifically truncated p95HER2. The p95HER2 fragment arises through alternative translation initiation or proteolytic cleavage of full-length HER2, yielding a membrane-bound protein that lacks the extracellular domain while retaining an active intracellular kinase domain. Because p95HER2 lacks the epitope bound by trastuzumab and other extracellular antibodies, it confers resistance to antibody-based therapeutics. However, because small-molecule inhibitors act intracellularly, Neratinib (HKI-272) directly engages the catalytic cleft of p95HER2. In the NALA trial biomarker analysis, tumors expressing p95HER2 exhibited improved PFS when treated with N+C compared to L+C, confirming that irreversible TKIs overcome trastuzumab resistance driven by extracellular receptor truncation.
Mutational profiling of pre-treatment tumors revealed distinct prognostic implications for somatic alterations in PIK3CA and HER2. Somatic mutations in the PIK3CA gene, which encode activating alterations in the p110α catalytic subunit of PI3K, were associated with reduced PFS across both the N+C and L+C treatment arms. Because PIK3CA mutations induce constitutive, receptor-independent activation of downstream AKT signaling, they mitigate the therapeutic benefit derived from upstream receptor inhibition. In contrast, somatic mutations within the HER2 kinase domain trended toward improved PFS in patients receiving N+C, suggesting that irreversible covalent engagement by neratinib maintains inhibitory efficacy against mutated HER2 conformations that might otherwise escape reversible inhibition.
These findings demonstrate that molecular characterization—incorporating HER2 protein expression levels, p95HER2 truncation status, and PIK3CA mutational profiling—provides critical prognostic and predictive insight for optimizing pan-HER inhibitor regimens in advanced HER2-positive metastatic breast cancer.
Biomarker Stratification and Therapeutic Efficacy of Covalent Pan-HER Inhibition
Copy code to add this image to your page
Neratinib (HKI-272) is an oral, irreversible pan-HER receptor tyrosine kinase inhibitor that covalently binds conserved cysteine residues in the ATP-binding pockets of HER2 (Cys-805), EGFR (Cys-797), and HER4. By permanently disabling transphosphorylation, it blocks downstream PI3K-AKT and MAPK proliferation signaling pathways. In metastatic breast cancer expressing high HER2 or truncated p95HER2 variants, covalent target engagement effectively prevents kinase reactivation, overcoming resistance mechanisms common to reversible TKIs or extracellular antibody therapies.
HER2 Protein Quantification Predicts Response to Irreversible Kinase Blockade
Quantitative measurement of HER2 protein density via immunohistochemical H-scoring and HERmark assays revealed a direct correlation between receptor abundance and progression-free survival outcomes. In the NALA trial cohort, tumors expressing high HER2 levels derived a pronounced benefit from combination therapy with Neratinib from Selleck compared to reversible inhibition with lapatinib. Downstream signaling reliance appears accentuated in tumors with dense receptor clusters. High HER2 expression amplifies dependence on the ErbB kinase cascade, rendering cancer cells exquisitely vulnerable to irreversible covalent inhibition. Receptor density remains a predictive indicator.
p95HER2 Expression and PIK3CA Mutational Status Modulate Clinical Progression
Truncated p95HER2 isoforms lack extracellular domain epitopes necessary for antibody binding, yet retain full intracellular catalytic capacity. Small-molecule TKIs penetrate the cell membrane. Experimental data showed that p95HER2-positive tumors retained sensitivity to covalent kinase blockade with Neratinib (HKI-272), yielding improved progression-free survival relative to lapatinib regimens. Conversely, somatic mutations in PIK3CA conferred inferior progression-free survival across all study cohorts. Activating PIK3CA mutations drive constitutive signaling downstream of the receptor level—effectively bypassing upstream HER2 inhibition. Target engagement was verified.